- LOGIN
- MemberShip
- 2026-10-13 11:13:02
- Hanmi's 'Epe' development team's 3-year focus…Behind Phase 3 success
- by Kim, Jin-Gu | translator Hong, Ji Yeon | 2026-10-13 08:45:34
"When we confirmed the statistical superiority result compared with a placebo group, we thought, 'This could be possible'"
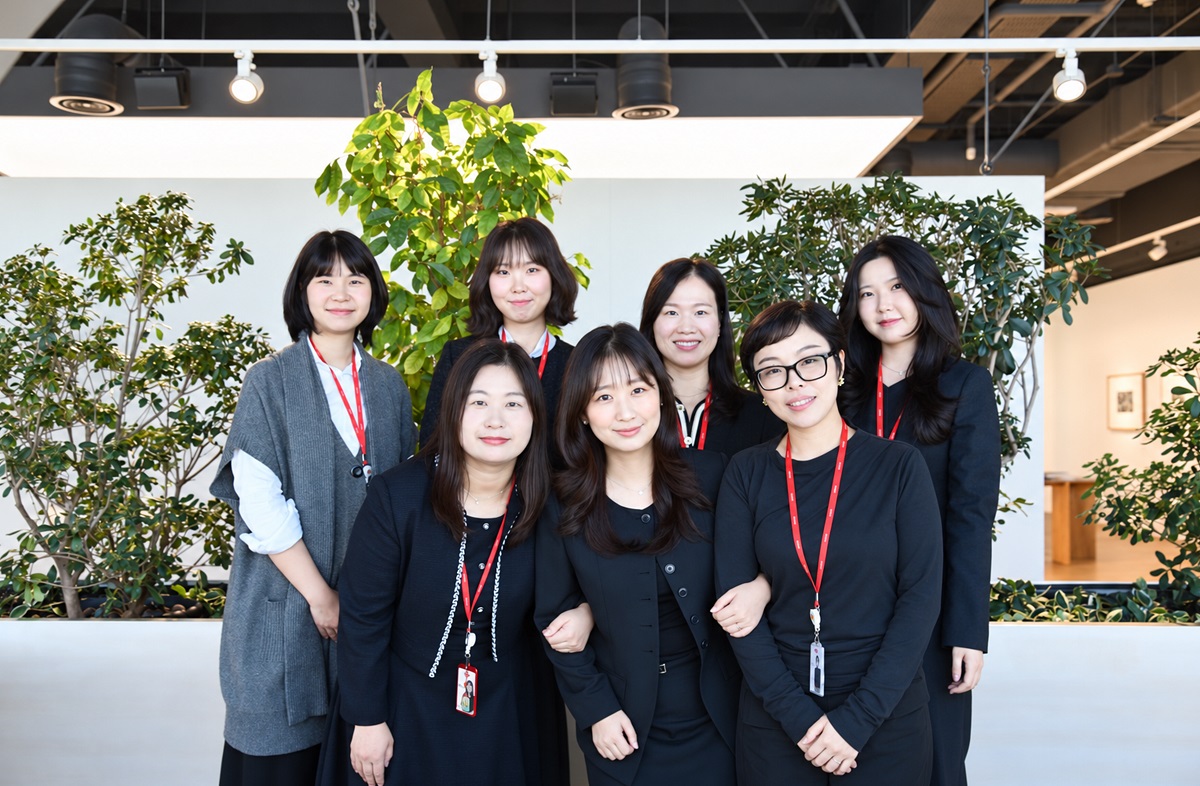
This was the moment when Group Leaders Kang Eun-sil and Jeong Min-sook, along with their team members in Hanmi Pharmaceutical's New Product Clinical Development Team, became convinced of the success of their novel obesity drug, 'Epe' (efpeglenatide). This happened when they first reviewed the internal readout of the Phase 3 clinical trial, conducted in 448 domestic patients.
The weight-loss efficacy was almost identical to what the development team had projected during trial design. Statistical superiority was demonstrated across both primary endpoints. It was the moment years of dedicated novel drug development were validated by the numbers.
However, after verifying the results, there was no time to celebrate. The team immediately had to prepare the clinical study report (CSR) and executive briefing materials. Responding to subsequent deficiency letters and information requests from the Ministry of Food and Drug Safety (MFDS) posed yet another critical hurdle.
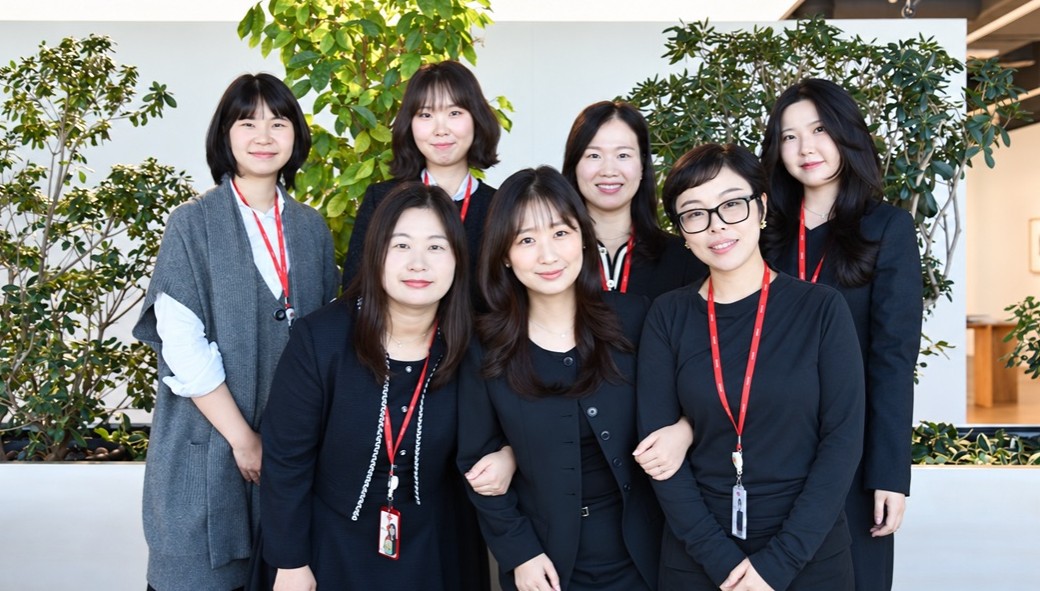
On the morning of the 7th, seven team members of Hanmi Pharmaceutical's New Product Clinical Development Team met to recall Epe's development journey. From trial design, patient recruitment, and investigative site management to data verification, statistical programming, and defending regulatory filings, each member faced distinct hurdles in their domain.
Just a few hours after the interview concluded, Epe received formal marketing authorization from the MFDS. It became the 45th domestically developed novel drug and the first homegrown GLP-1 receptor agonist obesity medication developed by a South Korean pharmaceutical company.
The MFDS approved five dose strengths: Epe Auto-Injector Inj 2·4·6·8·10 mg. In the domestic Phase 3 trial that served as the basis for approval, the mean body weight reduction at 40 weeks was 9.75% in the Epe group compared to 0.95% in the placebo group. The proportion of patients achieving at least a 5% reduction in body weight was 79.42% in the Epe group and 14.49% in the placebo group.
While the clinical outcomes are captured in concise figures, arriving at those data points was far from simple.
Relaunch after Sanofi's rights return… Past clinical data as an asset
The starting point for efpeglenatide was not as an anti-obesity medication, but as a treatment for type 2 diabetes. In 2015, Hanmi Pharmaceutical out-licensed efpeglenatide, formulated with its proprietary platform technology "LAPSCOVERY", to Sanofi. Although large-scale global clinical trials were subsequently conducted under Sanofi's lead, the rights were returned in 2020.
The event shook what had been a massive out-licensing achievement. Yet, Hanmi Pharmaceutical did not abandon development. Instead, the company uncovered new therapeutic potential within the accumulated clinical database. The decision to pivot the clinical strategy toward obesity crystallized around early 2023.
Kang Eun-sil, Head of the CR1 Group, explained, "We had secured a substantial amount of data from legacy Phase 1 and 2 trials as well as global diabetes clinical studies. We were able to leverage previously verified findings, such as pharmacokinetic (PK) and immunogenicity profiles, to design our obesity trial."
The legacy trial data proved particularly instrumental in determining the target maintenance dose and dose titration schedule. For obesity therapeutics, developers must achieve sufficient weight-loss efficacy while effectively managing early-phase gastrointestinal adverse events.
Lim Yi-rang, Section Manager of the CR1 Group, stated, "In previous trials, we had gathered extensive experience evaluating regimens ranging from non-escalating dosing schedules to dose-titration protocols adjusted every two or four weeks," adding, "Based on these data, we were able to establish an optimal target dose and titration algorithm tailored for patients with obesity."
This historical foundation also helped compress clinical timelines. Initiating a domestic Phase 3 trial required a rigorous scientific rationale for each design element, including target dosing and primary efficacy endpoints, and the pre-existing data package helped provide that justification.
Enrollment outpaced projections…Safety data capture tests clinical operations
In the domestic Phase 3 trial, faster-than-expected patient recruitment paradoxically added operational pressure to the team. Driven by intense public and clinical interest in obesity therapeutics, patient enrollment accelerated rapidly within a compressed timeframe. While rapid enrollment benefited the overall clinical development timeline, it substantially increased the workload for trial operations.
Jeong Min-sook, Head of the CR2 Group, explained, "Because a large number of subjects were enrolled in a short window, dosing initiation converged around the same timeframe," adding, "Consequently, the critical window requiring intensive safety monitoring overlapped, and subsequent end-of-treatment milestones converged as well."
The early dose-escalation phase was especially demanding. GLP-1 receptor agonists are prone to triggering gastrointestinal adverse events during early treatment, such as nausea, vomiting, and diarrhea. With a large cohort starting treatment at once, the volume of safety data that investigative sites had to collect surged.
Park Ji-young, Section Manager of the CO1 Group, recalled, "Because so many subjects enrolled over a short span, adverse events were gathered as well," adding, "Monitoring each trial center to ensure no safety reporting went omitted was an immense challenge."
Another operational hurdle was the pharmacokinetic evaluation. While legacy data from global diabetes trials were extensive, PK data in the Korean domestic population were lacking, requiring the integration of PK assessments directly into the Phase 3 protocol.
Park added, "Clinical investigators at Phase 3 study sites are primarily accustomed to assessing efficacy and safety endpoints," adding, "Managing them to execute blood draws and biospecimen collection protocols for PK evaluations, procedures typically handled in Phase 1 units, presented significant hurdles."
Verifying 448 patients' data in two months…The final database lock push
The final clinical visits of study participants also converged within the same window. The team had to clean, verify, and lock the clinical database for 448 subjects within a tight two-month window. Group Leader Jeong Min-sook recalled this phase as one of the program's most intense periods.
"The last subject completed their Week 40 visit in August, and the timeline required us to lock the database and review the results just two months later," Jeong recounted. "During those two months, we had to review every single data point across all 448 patients."
This went far beyond merely collating entered figures. It required reconciling body weight measurements, laboratory diagnostic panels, and adverse event narratives, followed by issuing and resolving site data queries for any missing or inconsistent values. Clinical trial managers monitored site operations, data management personnel verified data integrity, and medical monitors reviewed the dataset from a clinical perspective.
The statistical analysis was equally demanding. The development team adopted the estimand framework to prospectively define how intercurrent events, such as treatment discontinuation or the use of rescue anti-obesity medications, would be statistically accounted for.
Jang Jeong-joo, Section Manager of the DM/STAT Group, noted, "Managing a large dataset over an extended period was difficult, but implementing novel analytical methodologies was another major task," adding, "Drawing upon international clinical precedents and regulatory guidance, we established the statistical methodology and justified our rationale to the MFDS."
Clinical success followed by regulatory challenges… Months of MFDS deficiency inquiries
Despite meeting target endpoints in the Phase 3 trial, another formidable hurdle lay ahead: securing final marketing authorization. Kang Eun-sil identified the MFDS dossier review as the most grueling phase of the entire development program. While the clinical dossier had been compiled and submitted, it was met with an unprecedented volume of deficiency comments and supplementary inquiries.
"In all my years working in this field, it was the largest volume of supplementary review comments I have ever encountered," Kang recalled. "Typically, you receive one or two pages, but this time we received requests that were several times larger and called for exceptionally rigorous, extensive data packages."
The requests were equally demanding. Rather than merely supplying existing documentation, the team was required to run new subgroup analyses and substantiate the biological and clinical rationale of the findings. Kang added, "We spent countless hours deliberating whether to analyze data under this parameter or substantiate it via that methodology. Formulating the formal responses to the deficiency notices alone took several months."
This stringent regulatory review reflected Epe's status as the first domestically developed GLP-1 receptor agonist anti-obesity drug. Unlike foreign products being imported following prior global clearances, Epe was an originating product undergoing primary global approval review in South Korea.
The team explained that regulatory scrutiny focused on safety margins and the potential for off-label abuse. Kang remarked, "Given the public needs for cosmetic weight reduction, obesity therapeutics carry significant risks of off-label misuse," adding, "Consequently, the MFDS demanded in-depth safety datasets and adverse event characterizations."
In-house clinical execution without a CRO… Cross-functional synergy drives success
Throughout Epe's development, Hanmi Pharmaceutical avoided outsourcing core operations to contract research organizations (CROs), executing everything from clinical operations and data management to biostatistical programming. Although internal workloads were heavy, close cross-functional communication sped up troubleshooting and operational velocity.
Nam Soo-hee, a member of the CO1 Group, noted, "Managing the clinical trial directly without an external CRO allowed us to respond swiftly whenever issues arose. Instead of struggling in isolation, we could consult the project leader or data managers directly to reach immediate solutions."
For Lee Eun-hee, Senior Researcher in the DM/STAT Group, Epe was especially meaningful, as it was the first project assigned to her after joining the company. "My first assignment was a large-scale clinical trial carrying immense company expectations," adding, "Even for junior researchers, the team culture encouraged everyone to voice optimization ideas and collaborate on solutions."
Spanning more than three years, this development journey left the working-level team with value far beyond a completed clinical trial.
Jeong said, "Having worked on this program for over three years since early 2023, the entire team feels an attachment akin to raising a child. We hope Epe, built on so much dedication, achieves exceptional outcomes in the clinic." Lim also expressed, "Epe is a product forged through the blood, sweat, and tears of the clinical development team. Given the tremendous effort invested, we hope it reaches great heights."
After awaiting regulatory clearance until the morning of the 7th, the development team received news of final marketing authorization that afternoon. It culminated more than three years of effort, from protocol design through regulatory filing defense. The Epe development team's institutional knowledge is expected to serve as a platform for advancing future pipelines, including next-generation anti-obesity candidates.
Kang concluded, "Managing clinical trials across both obesity and diabetes in-house has yielded invaluable institutional expertise," adding, "Moving forward, our role is to leverage this execution capability to deliver robust clinical packages for upcoming pipeline candidates."
-
- 0
댓글 운영방식은
댓글은 실명게재와 익명게재 방식이 있으며, 실명은 이름과 아이디가 노출됩니다. 익명은 필명으로 등록 가능하며, 대댓글은 익명으로 등록 가능합니다.
댓글 노출방식은
댓글 명예자문위원(팜-코니언-필기모양 아이콘)으로 위촉된 데일리팜 회원의 댓글은 ‘게시판형 보기’와 ’펼쳐보기형’ 리스트에서 항상 최상단에 노출됩니다. 새로운 댓글을 올리는 일반회원은 ‘게시판형’과 ‘펼쳐보기형’ 모두 팜코니언 회원이 쓴 댓글의 하단에 실시간 노출됩니다.
댓글의 삭제 기준은
다음의 경우 사전 통보없이 삭제하고 아이디 이용정지 또는 영구 가입제한이 될 수도 있습니다.
-
저작권·인격권 등 타인의 권리를 침해하는 경우
상용 프로그램의 등록과 게재, 배포를 안내하는 게시물
타인 또는 제3자의 저작권 및 기타 권리를 침해한 내용을 담은 게시물
-
근거 없는 비방·명예를 훼손하는 게시물
특정 이용자 및 개인에 대한 인신 공격적인 내용의 글 및 직접적인 욕설이 사용된 경우
특정 지역 및 종교간의 감정대립을 조장하는 내용
사실 확인이 안된 소문을 유포 시키는 경우
욕설과 비어, 속어를 담은 내용
정당법 및 공직선거법, 관계 법령에 저촉되는 경우(선관위 요청 시 즉시 삭제)
특정 지역이나 단체를 비하하는 경우
특정인의 명예를 훼손하여 해당인이 삭제를 요청하는 경우
특정인의 개인정보(주민등록번호, 전화, 상세주소 등)를 무단으로 게시하는 경우
타인의 ID 혹은 닉네임을 도용하는 경우
-
게시판 특성상 제한되는 내용
서비스 주제와 맞지 않는 내용의 글을 게재한 경우
동일 내용의 연속 게재 및 여러 기사에 중복 게재한 경우
부분적으로 변경하여 반복 게재하는 경우도 포함
제목과 관련 없는 내용의 게시물, 제목과 본문이 무관한 경우
돈벌기 및 직·간접 상업적 목적의 내용이 포함된 게시물
게시물 읽기 유도 등을 위해 내용과 무관한 제목을 사용한 경우
-
수사기관 등의 공식적인 요청이 있는 경우
-
기타사항
각 서비스의 필요성에 따라 미리 공지한 경우
기타 법률에 저촉되는 정보 게재를 목적으로 할 경우
기타 원만한 운영을 위해 운영자가 필요하다고 판단되는 내용
-
사실 관계 확인 후 삭제
저작권자로부터 허락받지 않은 내용을 무단 게재, 복제, 배포하는 경우
타인의 초상권을 침해하거나 개인정보를 유출하는 경우
당사에 제공한 이용자의 정보가 허위인 경우 (타인의 ID, 비밀번호 도용 등)
※이상의 내용중 일부 사항에 적용될 경우 이용약관 및 관련 법률에 의해 제재를 받으실 수도 있으며, 민·형사상 처벌을 받을 수도 있습니다.
※위에 명시되지 않은 내용이더라도 불법적인 내용으로 판단되거나 데일리팜 서비스에 바람직하지 않다고 판단되는 경우는 선 조치 이후 본 관리 기준을 수정 공시하겠습니다.
※기타 문의 사항은 데일리팜 운영자에게 연락주십시오. 메일 주소는 dailypharm@dailypharm.com입니다.
- [Reporter’s View] Rebate investigation expertise must survive system overhaul
- Reporter's view | Kim, Jin-Gu
-
- "Besremi garners attention as a long-term treatment option for PV"
- Professor Dae-young Kim of the Department of Hematology-Oncology at CHA Bundang Medical Center